Lab Activities
Laboratory for Structure-based Molecular Design/ Drug Discovery Computational Chemistry Platform Unit
Research Activities
Team Director/ Unit Leader
Teruki Honma
To accelerate biological study and drug discovery, the aim of Laboratory for Structure-Based Molecular Design is to develop new technologies for in silico screening of biological tools and drug candidates. Our basic concept is the integration of both artificial intelligence (AI) and molecular simulations (molecular dynamics (MD), quantum mechanics (QM)) to design promising drug candidates for any target classes at any situations. PALLAS, MUSES, LAILAPS, QM-based model were developed and employedfor the efficient in silico screening based on the concept. Also, we constructed a large-scale small molecule database including more than 0.5 billion compounds that were preprocessed for various types of in silico screenings. In addition, we developed the integrated drug discovery AI platform collaborated with 17 pharmaceutical companies as well as Kyoto university sharing more than 15 million datapoints of assay data (AMED DAIIA).
The important mission of Drug Discovery Computational Chemistry Platform Unit is practical applications of the above-mentioned technologies developed by Laboratory for Structure-Based Molecular Design to drug discovery projects from academic and industrial research institutes. The tasks for the projects include hit identification, design for hit to lead and lead optimization by in silico screening. To handle the difficult tasks, we have daily built AI prediction models for Absorption, Distribution, Metabolism, Excretion, and Toxicity (ADMET) in addition to simulation-based approaches such as PALLAS, MUSES, and FMO method. The close collaborations with other platform units such as Protein Analysis Platform Unit (IMS, Dr. Shirouzu) and Medicinal Chemistry Platform Unit (CSRS, Dr. Koyama) are also very important to efficiently drive the projects by repeated cycles of design-synthesis-assay-protein structure analysis. We have been already involved in more than 40 drug discovery projects from RIKEN Program for Drug Discovery and Medical Technology Platforms (DMP) and AMED BINDS, P-PROMOTE as well as other academic research institutes and pharmaceutical companies.
Drug discovery AI platform developed by AMED DAIIA project
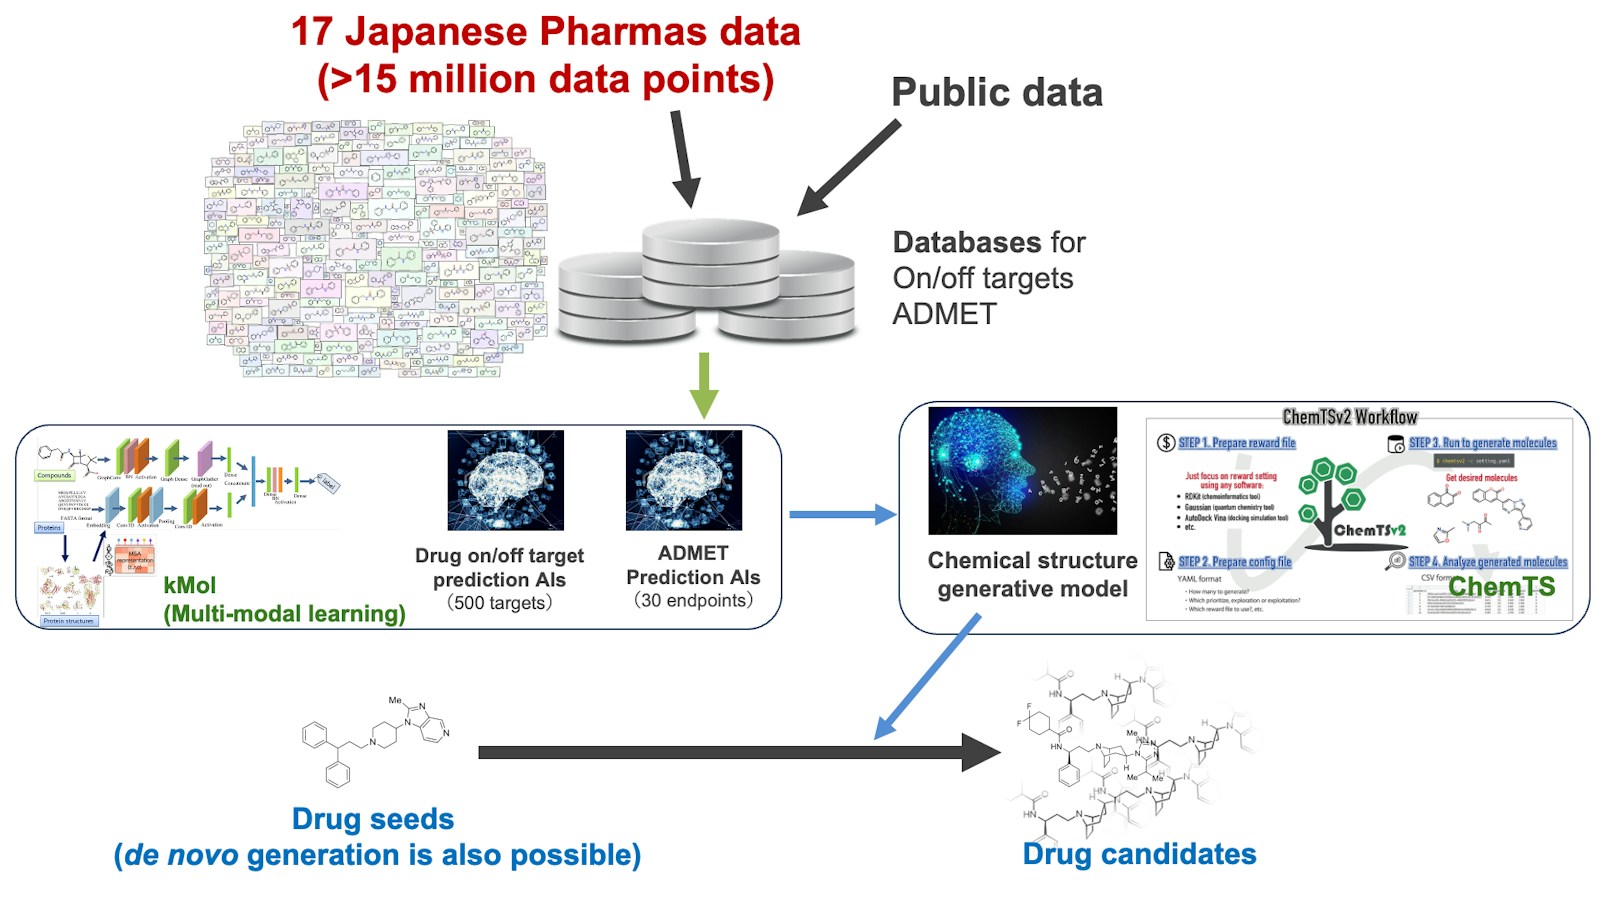
Using a dataset of over 15 million datapoints provided by 17 pharmaceutical companies, we developed AI prediction models for on/off-target effects and ADMET using multi-modal learning. By integrating these with structure generative models like ChemTS, the platform can efficiently identify and propose promising drug candidates.
Recent Major Publications
Kimishima A, Ikeda T, Takahashi O, Naher K, Watanabe C, Takai-Todaka R, Haga K, Honma S, Ujie Y, Uematsu T, Honsho M, Sunazuka T, Honma T, Katayama K, Asami Y. A Dual-Target-Based Screening Strategy for Anti-SARS-CoV-2 Active Compounds Enabling the Identification of Macrocyclic Peptide Natural Products: Chloropeptins. J Nat Prod 88, 2351–2359 (2025)
Ishida S, Sato T, Honma T, Terayama K. Large language models open new way of AI-assisted molecule design for chemists. J Cheminform 17, 36 (2025)
Watanabe C, Kamisaka K, Takaya D, Kato K, Tsuda K, Shu K, Murayama D, Isobe T, Ohyama T, Kato A, Fukuzawa K, Honma T. Developments and Activities in FMODB up to 2025: A Release Note. Chem-Bio Informatics Journal 25, 130–139 (2025)
Yagi Y, Kimura T, Watanabe C, Okiyama Y, Tanaka S, Honma T, Kaoru F. Comprehensive Protein-Ligand Interaction Analysis: Fragment Molecular Orbital Calculation on the Complexes of Human Protease Renin and its Inhibitors. Chem-Bio Informatics Journal 25, 107–129 (2025)
Masuda T, Watanabe C, Kato K, Honma T, Ohta M, Ikeguchi M. Quantitative Structure–Activity Relationships for Human Galectin-3 Inhibitors: Insights from Quantum Chemical Interaction Energy Terms. J Chem Inf Model 65, 6287–6297 (2025)
Nishigaya Y, Takase S, Sumiya T, Kikuzato K, Hiroyama T, Maemoto Y, Aoki K, Sato T, Niwa H, Sato S, Ihara K, Nakata A, Matsuoka S, Hashimoto N, Namie R, Honma T, Umehara T, Shirouzu M, Koyama H, Nakamura Y, Shirai F. Discovery of potent substrate-type lysine methyltransferase G9a inhibitors for the treatment of sickle cell disease. Eur J Med Chem 293, 117721 (2025)
Matsuoka S, Osada N, Kubota H, Kikuzato K, Koyama H, Sonoda T, Idei A, Yoshida M, Kikuchi M, Umehara T, Watanabe C, Honma T, Yasui H, Ikeda S, Takahashi N, Nakasone H, Kikuchi J, Furukawa Y. Discovery of a novel class NSD2 inhibitor for multiple myeloma with t (4; 14)+. Blood Neoplasia 2, 100091 (2025)
Yoshizawa T, Ishida S, Sato T, Ohta M, Honma T, Terayama K. Yoshizawa T, A data-driven generative strategy to avoid reward hacking in multi-objective molecular design. Nat Commun 16, 2409 (2025)
Akinaga Y, Terayama K, Kojima R, Harada Y, Takemura K, Honma T, Kitao A, Okuno Y. Precision spatiotemporal analysis of large-scale compound–protein interactions through molecular dynamics simulation. PNAS nexus 4, pgaf094 (2025)
Takayama K, Sato T, Honma T, Yoshida M, Inoue S. Inhibition of PSF activity overcomes resistance to treatment in cancers harboring mutant p53. Mol Cancer Ther 24, 370-383 (2025)
Invited Presentations
Teruki Honma. Development of an AI Platform for Drug Discovery and Its Applications. BDR-IPR Joint Symposium 2025, Osaka, Jan (2026)